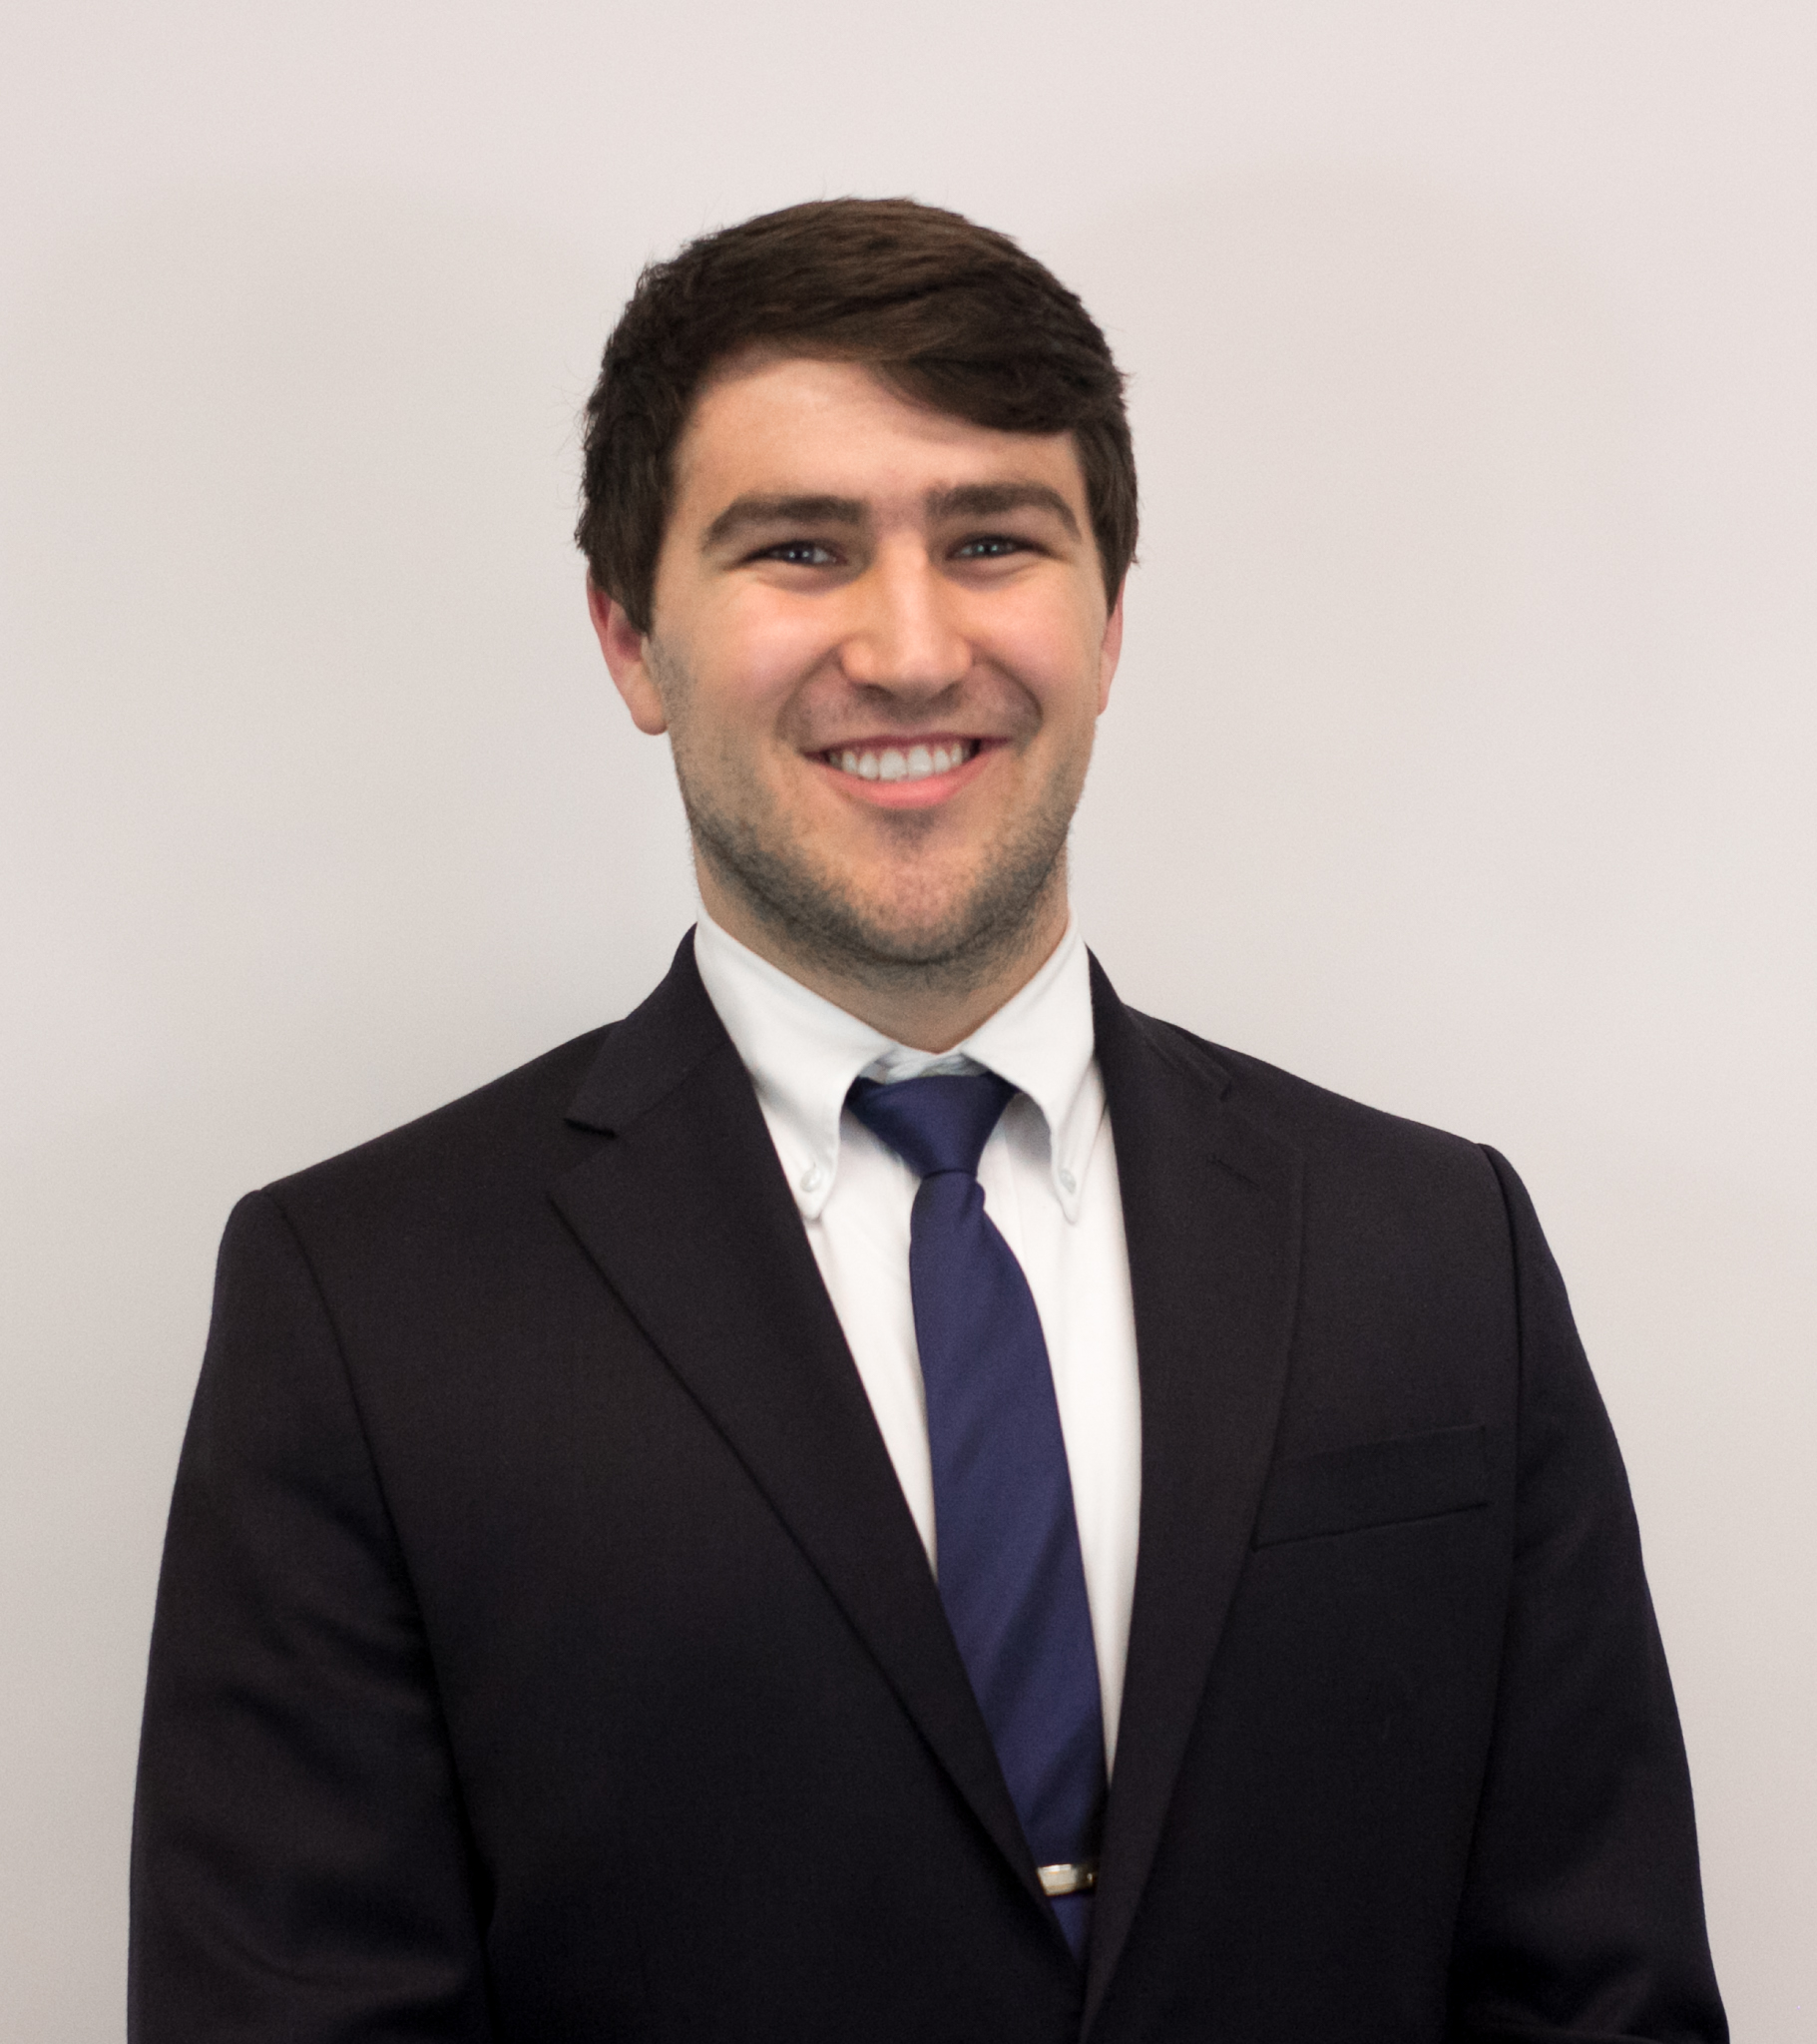
Alex Hoffman
My Background
My name is Alexander Hoffman. I’m a chemical engineer with expertise in computational heterogeneous catalysis, chemical kinetics, and machine learning. I earned my BS in Chemistry from the College of William and Mary in 2015, after which I worked at the EPA for a year. I realized that I wouldn’t be able to make it far without a graduate degree, so I moved to Gainesville, FL to attend the University of Florida. Initially, I intended only to get a MS in chemical engineering; however, I started doing research with Prof. David Hibbitts and enjoyed it so much that I stayed for 4 more years than I had planned, earning my Ph.D. under his guidance in 2022. During my Ph.D., I worked on a variety of projects with many excellent experimental groups, namely the Gounder group at Purdue University, the Schwartz group at University of Maine, and the Christopher group at UCSB. After my Ph.D., I moved to Cambridge, MA to do postdoctoral research with Prof. Rafa Gómez-Bombarelli. At MIT, I was lucky to work with Prof. Yuriy Román-Leshkov’s group and Dr. Manuel Moliner at ITQ in Valencia to study zeolite synthesis. In 2025, I moved to New York, NY to join Mirror Physics as a founding scientist.
When I’m not doing research, I enjoy cooking, playing sports—tennis, running, basketball, and swimming—watching classic films, reading, listening to music, and spending time with my wife, Becca, and dog, Bella.
My Research
I am always excited to discuss my work with others. Please feel free to reach out on LinkedIn if you cannot access any of the papers at the links below.
University of Florida
Brønsted Acid Zeolite Catalysts for Alcohol Dehydration
Much of my work during my Ph.D. focused on acid strength and alcohol dehydration chemistry in Brønsted acid-form zeolites. I used purely theoretical metrics like deprotonation energy (DPE) and experimentally measurable values like ammonia binding energies to assess how two nearby Brønsted acid sites interact in CHA zeolites and how sites on the surfaces of zeolites differ from those in the bulk in MFI. I also studied methanol and ethanol dehydration reactions on pairs of nearby Brønsted acid sites, where I found that nearby coadsorbed species can form hydrogen bonds with sites where reactions occur, and studied the effects of molecular clustering on methanol dehydration on single sites in CHA. These studies of methanol and ethanol dehydration were done in collaboration with Raj Gounder’s group. I also worked with a master’s student in the Hibbitts group, Tsung-wei Liu, and the Schwartz group to study hydroxymethyl furfural and ethanol cross-etherification in BEA zeolites. I was also momentarily preoccupied by issues with the default structure of MFI from the IZA database and how it can misbehave during some calculations. This paper (my first first-author paper) taught me the importance of using the right models for calculations in VASP!
Supported Rh Catalysts for Pollution Abatement
My second major project during my Ph.D. was on Rh catalysts for NOx reduction. I worked with the Christopher group at UCSB to characterize Rh nanoparticles and single-atoms on γ-Al2O3 supports. Rh can be particularly frustrating to work with because it reversibly aggregates to nanoparticles and disperses to single-atoms on oxide supports; as such, the active sites for NOx reduction can change over the course of the reaction. These projects sparked my interest in single-atom metal catalysts and the promise they provide for maximizing precious metal use and tailoring the behavior of active sites.
MIT
Controlling Zeolite Synthesis
I did some work studying zeolite synthesis during my Ph.D. in collaboration with the Gounder group—in particular, analyzing the way that Al is substituted near one another in MFI and which of those pairs are titrated by Co2+ and how tetrabutyl ammonium and tetrapropyl ammonium co-occlude with Na+ during MEL and MFI synthesis. However, my work during my Ph.D. used exclusively density functional theory (DFT) to characterize materials and study reaction mechanisms, but these tools can only model relatively small systems, relatively short time-scales, and are generally too slow to rapidly screen the full materials space that heterogeneous catalysts occupy. Therefore, I moved to MIT to work with Rafa Gómez-Bombarelli to learn more about machine learning (ML) methods as they pertain to zeolite synthesis and other computational chemistry challenges. During my postdoc, I developed an approach to learn new descriptors to predict zeolite synthesis outcomes and helped map the synthesis space of OSDAs for known zeolites to find cheaper synthesis routes for new compositions. I also contributed to DiffSyn, a generative diffusion model for planning materials syntheses, and to a workflow for high-throughput transition-state searches in zeolite nanopores.
Machine-Learned Interatomic Potentials
I also worked on EquiformerV3, an efficient and expressive SE(3)-equivariant graph attention transformer for predicting energies and forces of atomistic systems.
Mirror Physics
As a founding scientist at Mirror Physics, a start-up in New York, I worked across physics-based simulation, machine learning, and AI agents:
- Developed and tested new foundational machine-learned interatomic potential (MLIP) architectures for materials science and chemistry
- Helped build and test LLM-based agentic systems that run computer-assisted drug design (CADD) workflows, including DFT, classical molecular dynamics, docking, and free energy perturbation (FEP), for the company’s product, Axon
- Designed drug candidates using modern CADD tools, including docking, generative chemical models, free energy methods, and ML-based synthesis prediction
What I’m Looking For
I’m interested in industry roles that combine machine learning, atomistic simulation, and chemistry to design and understand materials and catalysts. I enjoy building computational tools that work closely with experimental teams, and I’m particularly excited about applications in sustainability, such as plastics and air pollution.
